

Peptide batch consistency lab quality control is the set of analytical and documentation practices that guarantee each peptide batch meets defined purity, identity, and stability standards before use in research. Without these controls, results across experiments become unreliable and regulatory submissions fall apart. Health Canada requires impurity profiles, forced degradation studies, and stability data aligned with ICH Q3A and Q3B guidelines. Analytical methods like HPLC and mass spectrometry form the technical backbone of any credible quality assurance programme. Peptilab supplies research-grade peptides with batch-specific certificates of analysis to support exactly this kind of rigorous lab work.
What are the essential analytical methods for peptide batch consistency?
HPLC and mass spectrometry answer two fundamentally different questions. HPLC measures how much of the target peptide is present relative to all detectable species. Mass spectrometry confirms what the molecule actually is by verifying molecular weight and fragmentation pattern.
Running only HPLC gives you a purity number without identity confirmation. That gap is a real risk. A peptide can show 98% purity by HPLC while still being the wrong sequence or a structurally similar impurity that co-elutes under your method conditions. Identity confirmation by mass spectrometry closes that gap.
LC-MS combines both techniques in a single run, delivering purity quantification and molecular identity simultaneously. For labs processing high volumes of batches, LC-MS reduces turnaround time without sacrificing analytical depth. It is the preferred approach when both speed and completeness matter.
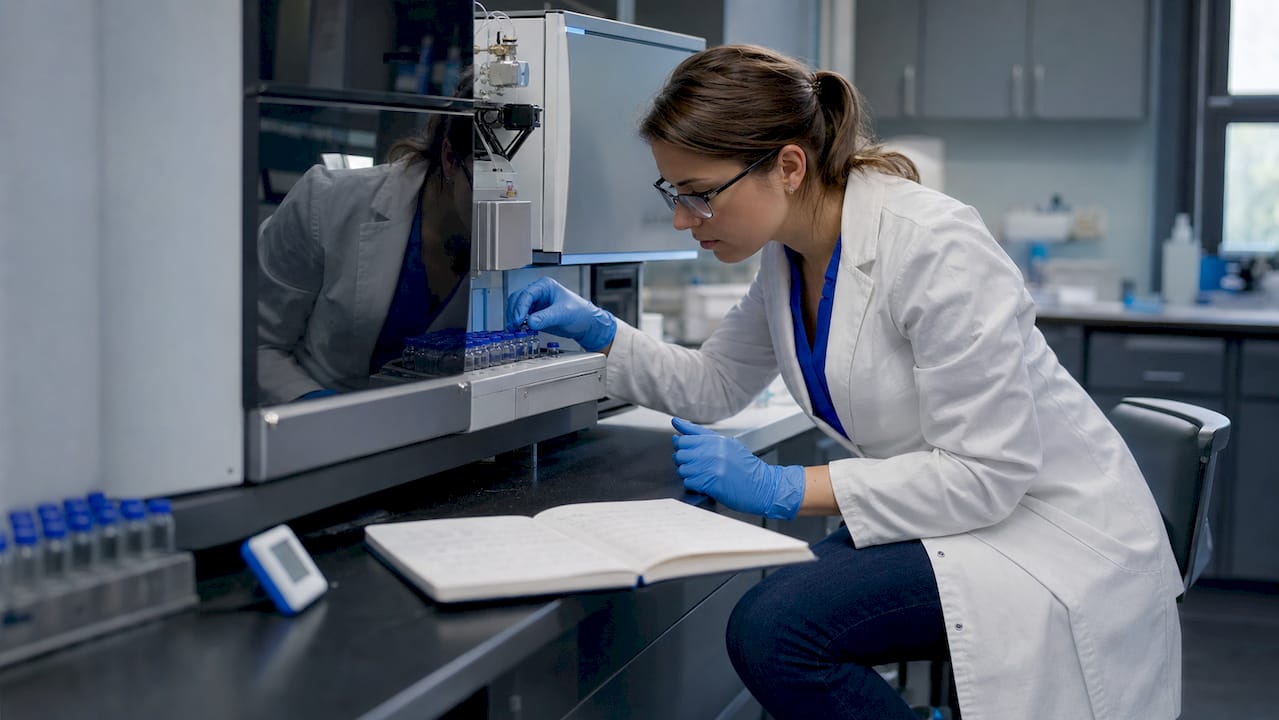
Pro Tip: When reviewing a COA, check whether purity was measured by HPLC-UV at 214 nm or 220 nm. Absorbance at 214 nm detects the peptide bond and gives a more representative purity value than 220 nm, which is more sensitive to aromatic residues and can skew results for certain sequences.
The table below compares the two primary methods across key evaluation criteria.
| Criterion | HPLC | Mass spectrometry |
|---|---|---|
| Primary output | Purity percentage | Molecular identity |
| What it measures | Relative peak area | Molecular weight and fragmentation |
| Detects wrong sequence | No | Yes |
| Detects co-eluting impurities | Partially | Yes, with MS/MS |
| Required for regulatory submission | Yes | Yes |
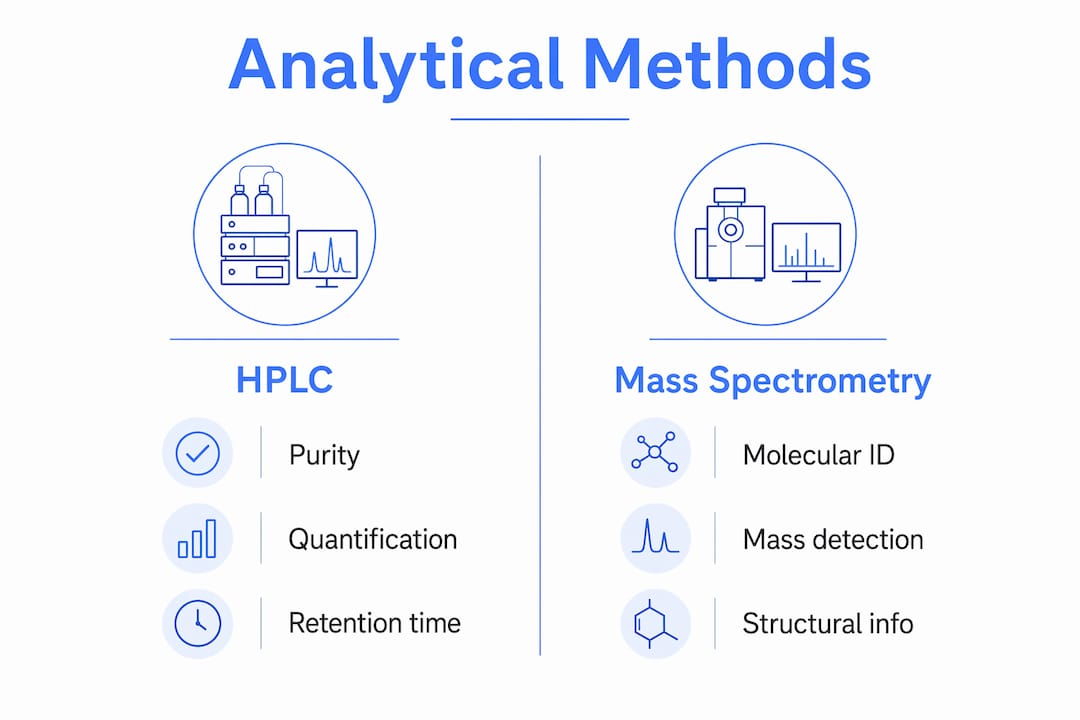
Pairing HPLC with mass spectrometry is the most reliable assurance of peptide purity and identity for quality control. Neither method alone satisfies the analytical expectations of Health Canada or the FDA for peptide sameness studies.
Which regulatory requirements govern peptide batch quality in Canadian labs?
Health Canada mandates analytical data packages that go well beyond a single purity percentage. Abbreviated New Drug Submission requirements include comparative impurity profiles, forced degradation studies covering acid, base, oxidation, heat, and light conditions, and stability data consistent with ICH Q3A and Q3B thresholds.
These requirements align closely with FDA expectations, which means labs preparing submissions for both regulators can build a single analytical package that satisfies both. The key is starting degradation and stability studies early in development. Stability of impurity profiles over time is a regulatory hurdle that catches many teams off guard when left too late.
A meaningful Certificate of Analysis does more than list a purity number. COAs that lack metadata such as method name, lot number, test date, instrument type, and integration parameters should be treated as administrative records rather than analytical proof. That distinction matters during an audit.
Key documentation elements that regulators expect to see:
- Impurity profile with identified and unidentified impurity thresholds per ICH Q3A/Q3B
- Forced degradation data covering all five stress conditions
- Stability data at defined time points and storage conditions
- COA metadata including method, lot number, test date, and analyst identifier
- Batch records linking each analytical result to a specific production lot
Forced degradation studies simulate real-world stress conditions to reveal how a peptide degrades and what impurities form. This data predicts shelf life and supports the impurity profile submitted to regulators. Labs that skip this step often face requests for additional data that delay approval timelines significantly.
How to implement batch documentation and chain-of-custody practices
A complete batch documentation template links every vial from purchase order through storage, aliquoting, use, and final disposition. This lifecycle record is what makes retrospective investigation possible when a result looks anomalous.
A practical batch record for each peptide lot should capture the following in order:
- Supplier information: Name, contact, and country of origin
- Order details: Purchase order number, order date, and expected delivery
- Receipt inspection: Condition on arrival, temperature log, and packaging integrity
- COA review: Purity, identity method, lot number, test date, and analyst
- Storage assignment: Location, temperature, and date placed in storage
- Aliquot log: Date, volume, concentration, and operator for each aliquot
- Use log: Experiment ID, date used, quantity, and operator
- Disposition: Quarantine, reject, or archive status with rationale
Chain-of-custody logs from receipt to disposition are critical for reconstructing batch history during troubleshooting or regulatory audits. A common error is saving only the COA PDF without capturing the surrounding data. The COA alone cannot tell an investigator who handled the vial, how it was stored, or whether a temperature excursion occurred.
Pro Tip: Build your batch record template in a structured spreadsheet or laboratory information management system with mandatory fields. Leaving fields optional guarantees they will be skipped under time pressure, which is exactly when documentation gaps cause the most damage.
Storing all batch-related data in a structured format, including COAs, storage records, and deviation notes, supports audit readiness and batch traceability. Assign a review status to each record: active, quarantine, rejected, or archived. That single field saves hours during an investigation.
What are common challenges in maintaining peptide batch consistency?
The most frequent error in peptide quality assurance is treating HPLC purity as a complete quality picture. HPLC confirms purity but cannot detect microbial contamination or endotoxin. Using HPLC data to answer questions about sterility is a category error that puts downstream experiments and, in some contexts, study subjects at risk.
Purity numbers themselves require careful interpretation. Purity percentages on COAs depend on baseline construction, peak detection sensitivity, and run length. Two labs running the same peptide under different chromatographic conditions can report meaningfully different purity values. QC professionals must review the method specifics rather than accept numbers at face value.
“A 98% purity result is only as meaningful as the method that produced it. Without knowing the column type, gradient, detection wavelength, and integration parameters, that number is a starting point for questions, not a conclusion.”
Stability-related degradation is another source of batch inconsistency that documentation alone cannot prevent. Peptides stored above their recommended temperature, exposed to freeze-thaw cycles, or reconstituted in incompatible solvents will show impurity profile drift over time. Retesting schedules tied to storage duration and conditions catch this drift before it contaminates experimental results.
Practical steps to reduce batch consistency failures:
- Confirm identity by mass spectrometry on every new lot, not just purity by HPLC
- Run separate validated microbiological assays for sterility and endotoxin when required
- Review COA method details before accepting a purity number as valid
- Set retesting intervals based on stability data, not arbitrary calendar dates
- Log every freeze-thaw cycle and temperature excursion in the batch record
Key takeaways
Reliable peptide batch quality requires combining HPLC purity data with mass spectrometry identity confirmation, maintaining complete chain-of-custody documentation, and meeting Health Canada’s ICH-aligned regulatory standards from the earliest stages of development.
| Point | Details |
|---|---|
| Use HPLC and MS together | Purity alone does not confirm identity; both methods are required for complete batch assessment. |
| Meet Health Canada standards | Submit impurity profiles, forced degradation data, and stability results per ICH Q3A/Q3B. |
| Build complete batch records | Capture supplier info, COA metadata, storage logs, aliquot history, and disposition status. |
| Interpret COAs critically | Review method details, not just the purity number, before accepting a batch as compliant. |
| Retest on a stability schedule | Tie retesting intervals to actual stability data to catch impurity drift before it affects results. |
What I have learned from years of watching COAs get misread
The most persistent problem in peptide quality control is not a lack of data. It is misplaced confidence in incomplete data. QC professionals receive a COA showing 99% purity and treat the batch as fully characterised. That purity number reflects one method, one instrument, and one set of integration parameters. It says nothing about sequence accuracy, endotoxin load, or what happens to the impurity profile after six months at minus 20°C.
Purity results influenced by chromatographic parameters are a real and underappreciated variable. I have seen two COAs for nominally identical peptides from different suppliers show a 3% purity difference that disappeared entirely when both were run on the same column under the same gradient. The peptides were equivalent. The methods were not.
The labs that handle this well share one habit: they treat every new lot as a question to be answered, not a box to be ticked. They pull the mass spec data alongside the HPLC trace. They check the test date against the stability window. They log the receipt inspection before the vial goes into the freezer. That culture of documentation does not slow research down. It prevents the kind of batch-related failures that cost weeks of repeated experiments.
The regulatory environment is also tightening. Health Canada’s alignment with FDA and ICH standards means the analytical package that satisfied a submission three years ago may not satisfy one today. Starting forced degradation and stability studies early is no longer optional for teams working toward regulatory approval. It is the baseline expectation.
— Admin
Peptilab’s research peptides for quality-controlled labs
Quality control professionals need suppliers who match their documentation standards, not just their purity targets.
Peptilab supplies research-grade peptides with batch-specific COAs that include lot numbers, test dates, and analytical method details, giving your team the metadata needed for compliant batch records. Every product is verified through third-party testing with purity greater than 99%. Peptilab also supports Canadian regulatory alignment, with domestic fulfilment that eliminates import delays and keeps your supply chain predictable. For labs working across metabolic, cellular, or recovery research, the full peptide catalogue covers a broad range of characterised compounds backed by transparent analytical data.
FAQ
What is peptide batch consistency in lab quality control?
Peptide batch consistency is the degree to which successive production lots of a peptide meet the same defined standards for purity, identity, and impurity profile. Lab quality control confirms this consistency through analytical testing and documented batch records.
Why is HPLC alone not enough for peptide quality assurance?
HPLC measures relative purity but cannot confirm molecular identity or detect microbial contamination. Mass spectrometry is required alongside HPLC to verify the peptide sequence and molecular weight.
What does Health Canada require for peptide batch documentation?
Health Canada expects impurity profiles, forced degradation data covering acid, base, oxidation, heat, and light conditions, and stability data consistent with ICH Q3A and Q3B thresholds as part of a complete analytical submission package.
How often should peptide batches be retested for stability?
Retesting intervals should be set based on the peptide’s stability data rather than fixed calendar dates. Forced degradation studies define the expected impurity profile changes over time and should guide your retesting schedule.
What makes a COA analytically valid rather than just administrative?
A valid COA includes the analytical method name, instrument type, lot number, test date, analyst identifier, and integration parameters. COAs missing this metadata are administrative records and cannot be used as analytical proof of batch quality.