

Peptide chromatography is the separation and analysis of peptides in a mixture using liquid chromatography to assess purity, identify impurities, and confirm molecular identity. The dominant technique is reversed-phase HPLC (RP-HPLC), which separates peptides by hydrophobicity and detects them at 214 nm, targeting an area under the curve (AUC) purity threshold of at least 98%. Understanding peptide chromatography basics is not optional for researchers working with synthetic or recombinant peptides. A single co-eluting impurity can invalidate a bioassay, skew dose-response data, or trigger a regulatory rejection. This article covers how the technique works, which methods suit which applications, and how to avoid the most common analytical errors.
What is peptide chromatography explained: principles and techniques
Peptide chromatography works by exploiting differences in how individual peptides interact with a stationary phase inside a chromatographic column. In RP-HPLC, the stationary phase is hydrophobic and the mobile phase is aqueous, so peptides elute in order of increasing hydrophobicity as the organic solvent concentration rises during a gradient run. The peptide bond absorbs UV light strongly at 214 nm, making this wavelength the standard detection point for universal peptide quantification.
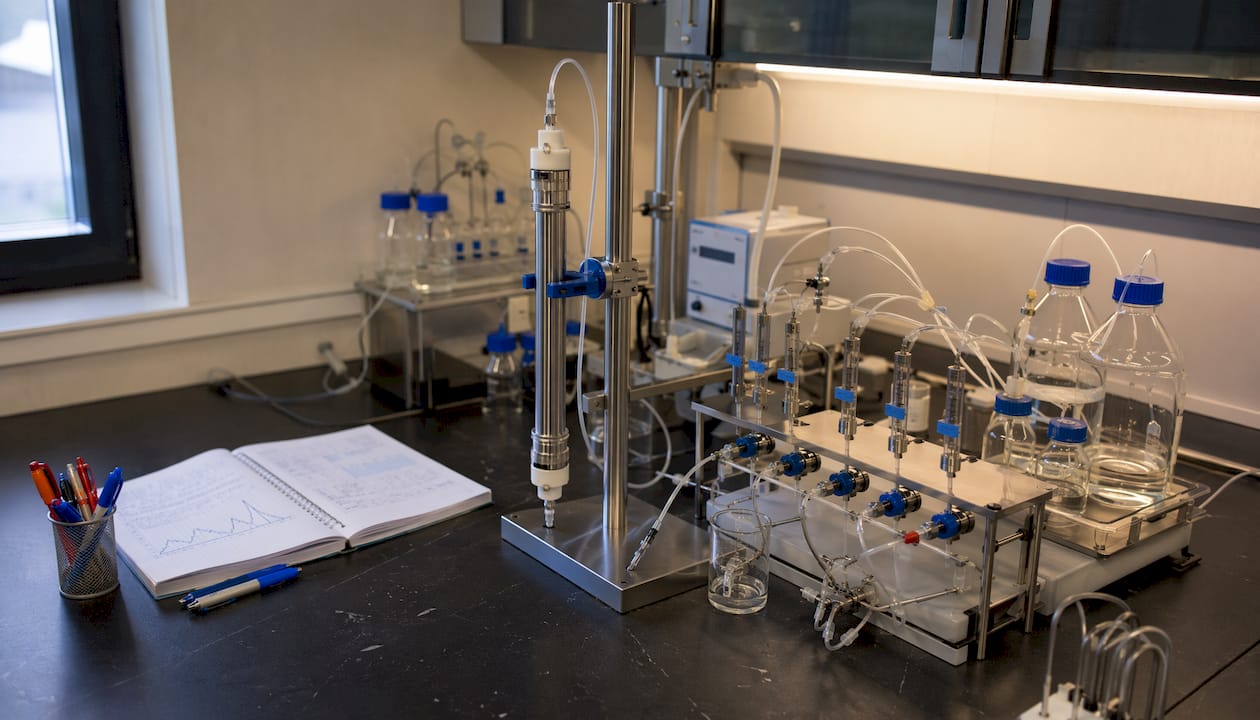
Mobile phase composition directly controls resolution and peak shape. 0.1% trifluoroacetic acid (TFA) is the most widely used ion-pairing reagent in RP-HPLC because it suppresses ionisation of basic residues, sharpens peaks, and improves retention reproducibility. Without an ion-pairing agent, basic peptides produce broad, asymmetric peaks that inflate apparent purity values.
Column chemistry selection follows peptide size and hydrophobicity:
- C18 columns suit short peptides (fewer than 15 residues) and moderately hydrophobic sequences.
- C8 columns offer reduced retention for mid-length peptides and help avoid irreversible adsorption.
- C4 columns are the standard choice for large or highly hydrophobic peptides, including many therapeutic candidates.
- 300 Å pore size is recommended for peptides over 30 residues to allow full access to the stationary phase surface.
Single-dimension chromatography has a hard limit. HPLC alone cannot distinguish co-eluting species with similar hydrophobicity, which means a peptide can appear 99% pure by UV while carrying a structurally related impurity at a biologically relevant level. Coupling RP-HPLC with mass spectrometry resolves this problem by adding a molecular weight dimension to the purity assessment.
Pro Tip: Run your peptide on two different column chemistries, such as C18 and C8, before reporting a final purity value. A peptide that reads 98% on C18 may show a distinct impurity shoulder on C8 due to co-elution phenomena on the first column.
How does advanced peptide chromatography handle impurity profiling?
Single-dimension RP-HPLC misses impurities that co-elute with the target peptide under a single set of conditions. Two-dimensional liquid chromatography (2D-LC) solves this by coupling two columns operating at orthogonal pH values, so an impurity that co-elutes at low pH separates cleanly at high pH. pH-orthogonal 2D-LC increases separation coverage by approximately 68% over single-dimensional methods. That gain translates directly into more complete impurity profiles and fewer false-clean results.
Isobaric impurities present a particular challenge. Two peptides can share the same nominal molecular weight yet differ in sequence, deamidation state, or stereochemistry. High-resolution mass spectrometry, such as Orbitrap or Q-TOF platforms, resolves these species at the sub-dalton level when combined with chromatographic separation. Neither technique alone is sufficient for confident impurity assignment.
Regulatory requirements reinforce this multi-method approach. ICH Q6B mandates multi-enzyme digestion workflows for peptide mapping, requiring enzymes such as trypsin, Lys-C, and Glu-C to achieve complete sequence coverage. Single-enzyme digestion routinely misses Proline-rich regions, leaving gaps in the sequence map that regulators at Health Canada and the FDA will flag during biologic submissions.
| Method | Separation basis | Best application |
|---|---|---|
| RP-HPLC (C18) | Hydrophobicity | Routine purity screening, short peptides |
| RP-HPLC (C4, 300 Å) | Hydrophobicity | Large or hydrophobic peptides |
| pH-orthogonal 2D-LC | Hydrophobicity + charge | Comprehensive impurity profiling |
| Ion-pair RP + SAX-HPLC | Hydrophobic + ionic | Peptide-oligonucleotide conjugates |
| LC-MS (high-resolution) | Mass + retention time | Isobaric impurity identification |
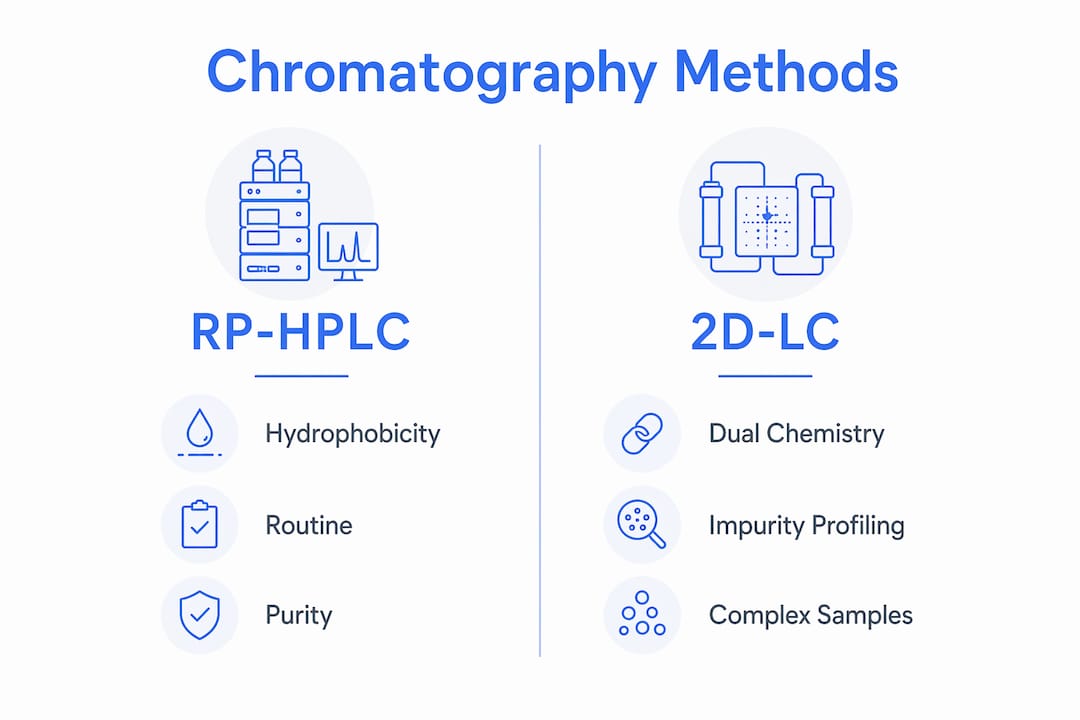
Pro Tip: When profiling a new synthetic peptide, run the 2D-LC screen before committing to a single-dimension method. Discovering a co-eluting impurity after scale-up is far more costly than catching it at the analytical stage.
Selecting and optimising chromatographic conditions for peptides
Stationary phase selection is the single most consequential method decision. Choosing C8 or C4 with 300 Å pores for peptides exceeding 30 residues prevents irreversible adsorption and the peak broadening that follows. Researchers who default to C18 for all peptides routinely report artificially low recoveries for larger sequences because the peptide binds too tightly to the stationary phase.
Mobile phase pH and acidic modifiers shape both retention and peak symmetry. TFA at 0.1% remains the standard, but formic acid is a viable substitute when downstream mass spectrometry is required, since TFA suppresses electrospray ionisation signal. The choice between these two modifiers depends on whether the primary goal is chromatographic resolution or MS sensitivity.
Temperature and gradient profile affect peak width and run time simultaneously. Higher column temperatures reduce mobile phase viscosity and improve mass transfer, producing sharper peaks. Shallow gradients increase resolution between closely eluting species but extend run time. The practical balance depends on sample throughput requirements and the complexity of the peptide mixture.
Baseline integrity is a direct indicator of data quality. Artificially flat chromatogram baselines are a red flag for digitally manipulated data, whereas minor, consistent electronic noise confirms a legitimate measurement. Researchers reviewing third-party certificates of analysis (COAs) should treat suspiciously smooth baselines as grounds for requesting raw data files.
Key method optimisation checkpoints:
- Confirm column pore size matches peptide molecular weight before the first injection.
- Verify TFA or formic acid concentration is consistent across all runs in a study.
- Set gradient slope to resolve the main peak from its nearest neighbour by at least 1.5 resolution units.
- Record column temperature and hold it constant across all replicates.
- Inspect baseline noise in every chromatogram before accepting a purity value.
What are the practical applications of peptide chromatography in research?
Peptide chromatography is the primary quality control tool for verifying peptide synthesis accuracy and detecting synthesis by-products such as deletion sequences, truncations, and oxidised residues. A synthesis run that looks clean by mass alone can carry deletion impurities at 2–5% that only chromatographic separation reveals. Catching these at the QC stage prevents downstream experimental failures.
Purity benchmarks are not arbitrary. Research-grade peptides require a minimum of 98% chromatographic purity for most biochemical assays, including receptor binding studies, enzyme kinetics, and cell-based models. Below this threshold, impurities can act as agonists, antagonists, or enzyme inhibitors in their own right, confounding results in ways that are difficult to trace retrospectively.
Mass spectrometry integration adds molecular weight confirmation to the purity picture. The combination of RP-HPLC retention time and accurate mass provides two independent lines of evidence for peptide identity, which is the minimum standard for peptide API sourcing in pharmaceutical development. Retention time alone is insufficient because structurally distinct peptides can share similar hydrophobicity values.
Peptide-oligonucleotide conjugates require a different analytical strategy entirely. Orthogonal chromatographic methods, specifically ion-pair reversed-phase HPLC and strong anion-exchange HPLC (SAX-HPLC), separate these conjugates based on both hydrophobic and charge characteristics. No single column chemistry resolves the full complexity of these hybrid molecules.
High-throughput applications have accelerated significantly with UHPLC. Dual-pump UHPLC workflows achieve a 10-fold increase in throughput, reducing injection-to-injection times from 123 minutes to 12.5 minutes while maintaining retention time control. For pharmaceutical peptide development programmes running hundreds of variants, this throughput gain is the difference between a week of instrument time and a single day.
Key takeaways
Peptide chromatography requires combining RP-HPLC with mass spectrometry, appropriate column chemistry, and rigorous baseline assessment to produce reliable purity and identity data.
| Point | Details |
|---|---|
| 98% purity is the minimum | Research-grade peptides must meet at least 98% chromatographic purity for valid biochemical assays. |
| Column chemistry determines outcome | Match stationary phase (C18, C8, C4) and pore size to peptide size and hydrophobicity to prevent adsorption. |
| 2D-LC closes coverage gaps | pH-orthogonal two-dimensional chromatography increases separation coverage by approximately 68% over single-dimension methods. |
| MS coupling is non-negotiable | RP-HPLC alone cannot distinguish co-eluting isobaric impurities; mass spectrometry adds the required second dimension. |
| Baseline noise confirms data integrity | Smooth, flat baselines in chromatograms indicate potential data manipulation; consistent electronic noise is the expected norm. |
Why I think researchers underestimate column selection
After working through peptide characterisation workflows across a range of molecular weights and sequence types, the single most common error I see is defaulting to C18 for every peptide. Researchers treat it as the universal column, and it is not. A 40-residue hydrophobic peptide on a C18 column with 100 Å pores will adsorb irreversibly, and the researcher will spend days troubleshooting recovery before realising the column was wrong from the start.
The second underestimated issue is the gap between chromatographic purity and actual peptide quality. A 98% AUC value is only as meaningful as the method used to generate it. Purity values vary significantly between C18 and C8 columns due to co-elution, which means a purity number without a stated method is nearly uninterpretable. Regulatory reviewers know this. Researchers presenting purity data without specifying column chemistry and gradient conditions are presenting incomplete data, full stop.
The future of this field is clearly moving toward UHPLC with multi-column configurations and integrated MS detection as the default, not the premium option. The throughput gains are too large to ignore, and the regulatory pressure for comprehensive impurity characterisation is only increasing. Researchers who build these methods now will be ahead of the curve when ICH guidelines tighten further.
— Admin
Research-grade peptides for Canadian scientists
Canadian researchers working with peptide chromatography need a supply source that matches the analytical rigour their methods demand.
Peptilab supplies research-grade peptides verified to greater than 99% purity, with certificates of analysis and third-party testing documentation included with every order. The catalogue covers peptides suited to metabolic research, receptor binding studies, and biochemical characterisation workflows. Domestic Canadian fulfilment means no import delays and no customs uncertainty. Researchers who need analytical documentation aligned with ICH Q6B expectations will find Peptilab’s peptide types and specifications a practical starting point for sourcing decisions.
FAQ
What is peptide chromatography used for?
Peptide chromatography separates peptides in a mixture to assess purity, identify impurities, and confirm molecular identity. It is the primary quality control method for synthetic and recombinant peptides in biochemical research.
What detection wavelength does peptide RP-HPLC use?
RP-HPLC detects peptides at 214 nm, where the peptide bond absorbs UV light strongly. This wavelength provides near-universal detection across all peptide sequences regardless of side-chain composition.
Why is 98% purity the standard for research peptides?
At purity levels below 98%, co-purified impurities can act as independent biological agents in assays, producing confounded results in receptor binding, enzyme kinetics, and cell-based studies. The 98% threshold is the accepted minimum for data reliability.
When should researchers use 2D-LC instead of standard RP-HPLC?
Two-dimensional LC is warranted when a peptide mixture contains structurally similar impurities that co-elute under a single set of chromatographic conditions. It increases separation coverage by approximately 68% and is particularly valuable for complex therapeutic peptide characterisation.
How does column choice affect reported peptide purity?
A peptide’s apparent purity can differ substantially between C18 and C8 columns because co-elution patterns change with stationary phase chemistry. Purity values must always be reported alongside the specific column chemistry and gradient conditions used to be scientifically interpretable.