

Peptide sequence verification is defined as the analytical confirmation that a peptide’s amino acid order matches its intended sequence, providing a molecular fingerprint that validates identity before any research application. In life sciences, this process is the foundation of research integrity. Without it, a peptide that appears pure by HPLC may still carry the wrong sequence, making every downstream result unreliable. The industry standard for this confirmation is tandem mass spectrometry (MS/MS), which fragments peptides under 50 residues to generate sequence-specific ion series. Regulatory frameworks such as ICH Q6A and supporting documentation like Certificates of Analysis (COAs) formalize these requirements across pharmaceutical and research settings.
What is peptide sequence verification and why does it matter?
Peptide sequence verification is the process of analytically confirming that each amino acid in a peptide occupies the correct position in the chain. A peptide with a single transposed or substituted residue can have identical molecular weight to the correct compound yet behave entirely differently in a biological assay. That distinction is invisible to HPLC purity data alone.
Peptide verification confirms identity, purity, sequence, and quantity to the standards required for research reproducibility and regulatory submission. This means a researcher relying solely on a purity percentage is working with incomplete information. Sequence data closes that gap.
The importance of peptide verification extends beyond individual experiments. In therapeutic development, a misidentified peptide can invalidate an entire study, delay regulatory submissions, or, in clinical contexts, create safety concerns. Sequence confirmation at the point of procurement is the first line of defense against these outcomes.
What are the primary techniques used for peptide sequence verification?
Tandem mass spectrometry (MS/MS)
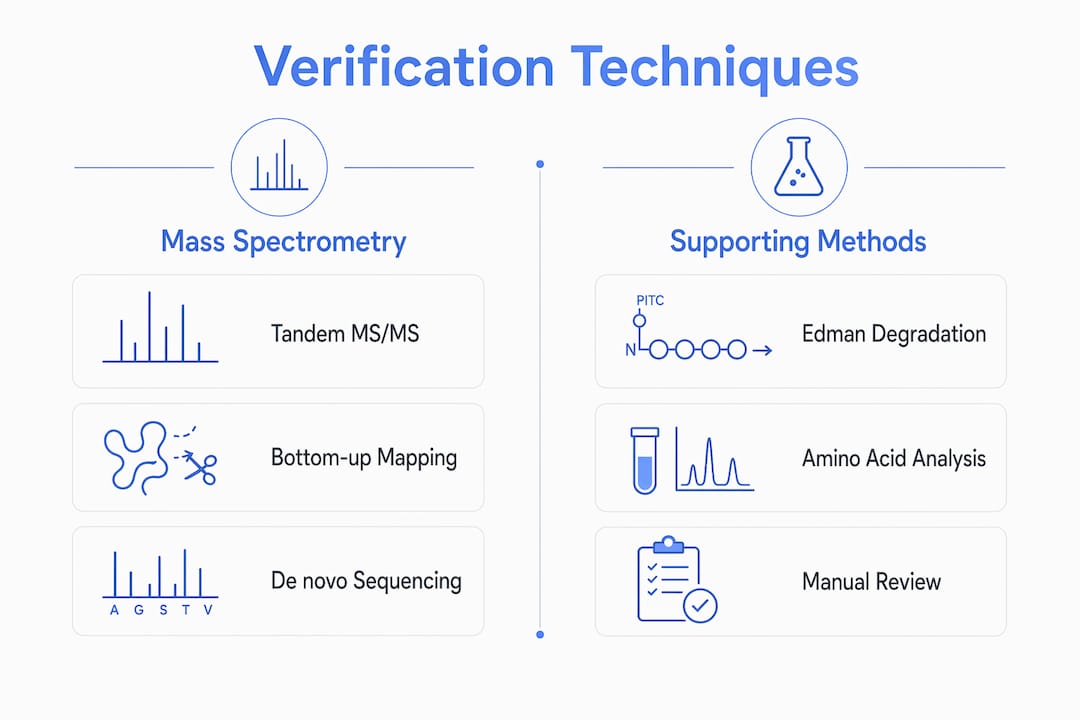
Tandem MS is the definitive method for peptide sequence analysis. The technique fragments a peptide at its peptide bonds, generating b-ions and y-ions that map directly to the amino acid sequence. Each fragmentation pattern is unique to a given sequence, functioning as a molecular fingerprint. MS/MS fragmentation is the accepted standard for synthetic peptides under 50 residues.
The strength of MS/MS lies in its specificity. Two peptides with identical molecular weight but different sequences produce distinct fragmentation patterns. No other single technique provides that level of discrimination at this throughput.
Bottom-up peptide mapping
Bottom-up peptide mapping uses enzymatic digestion, chromatographic separation, and MS/MS analysis to confirm the amino acid sequence of larger therapeutic proteins. The protein is digested with a protease such as trypsin, producing smaller peptide fragments that are individually sequenced. This method is an established standard for batch release and product characterization in biopharmaceutical manufacturing.
Peptide mapping is particularly valuable for monoclonal antibodies and other complex biologics where direct sequencing of the intact molecule is not practical. The method generates high sequence coverage and can detect post-translational modifications, oxidation artifacts, and deamidation events that affect product quality.
Supporting methods: Edman degradation and amino acid analysis
Edman degradation sequences a peptide from its N-terminus, cleaving one residue at a time for identification. The method is reliable for short peptides but becomes impractical beyond 30–50 residues and cannot sequence cyclic or N-terminally blocked peptides. Amino acid analysis quantifies the composition of residues after complete hydrolysis but does not confirm their order. Both methods serve as useful complements to MS/MS rather than replacements.
De novo peptide sequencing with advanced computational tools, including deep learning and mirror protease methods, improves coverage and confidence for novel peptides not present in reference databases. This approach is gaining traction for non-standard or modified peptides where database matching is insufficient.
| Method | Best application | Key limitation |
|---|---|---|
| Tandem MS/MS | Synthetic peptides under 50 residues | Requires high-quality spectra and expert review |
| Bottom-up peptide mapping | Therapeutic proteins and biologics | Labor-intensive sample preparation |
| Edman degradation | Short, unmodified peptides | Cannot sequence blocked N-termini |
| Amino acid analysis | Composition confirmation | Does not confirm sequence order |
| De novo sequencing | Novel or modified peptides | Computationally intensive |
Pro Tip: Always request both the MS/MS spectrum and the sequence assignment report from your supplier. A purity figure without fragmentation data does not constitute sequence verification.
How do researchers interpret peptide sequence data and avoid common pitfalls?
Automated software for peptide mapping has real limitations. Manual expert review is essential to avoid misassignments and artifacts that software alone cannot resolve. Researchers who rely entirely on automated outputs risk accepting incorrect sequence calls without realizing it.
The most common source of error is isobaric interference. Isobaric dipeptides such as SA vs. GT have identical nominal masses and can confuse mapping software, leading to incorrect sequence assignments. Only manual inspection of the MS/MS spectrum, combined with knowledge of the expected sequence, can resolve these ambiguities reliably.
Retention time is another misunderstood metric. Retention time alone cannot confirm peptide identity because deletion variants with similar hydrophobicity may co-elute under standard chromatographic conditions. MS/MS fragmentation patterns provide the definitive confirmation that chromatography cannot.
Best practices for accurate sequence validation include:
- Cross-reference every software-generated sequence assignment against the theoretical fragmentation pattern for the expected peptide.
- Flag any spectrum where fewer than 70% of expected b-ions or y-ions are matched.
- Treat abundant modifications such as oxidized methionine as potential artifacts before accepting them as real modifications.
- Use validated commercial software in regulated environments, but always follow with manual review by an experienced analyst.
Pro Tip: When working with peptides containing leucine and isoleucine, note that standard MS/MS cannot distinguish between these two residues due to identical masses. Confirm via Edman degradation or specialized ion mobility methods if the distinction is critical to your research.
What regulatory standards and documentation support peptide sequence verification?
The Certificate of Analysis is the primary document linking analytical evidence to a specific peptide batch. A valid COA does more than state a purity percentage. A complete COA must include identity, purity, sequence, and quantity, supported by traceable analytical evidence such as MS spectra and HPLC chromatograms.
ICH Q6A, the International Council for Harmonisation guideline on test procedures and acceptance criteria for new drug substances, defines the analytical requirements that apply to peptide-based active pharmaceutical ingredients. Under ICH Q6A, identity testing must use a method capable of distinguishing the compound from closely related substances. MS/MS satisfies this requirement. HPLC retention time alone does not.
A complete, research-grade COA includes the following elements:
- Peptide identity confirmed by MS/MS fragmentation data, not just molecular weight.
- Purity expressed as a percentage by HPLC area, with the chromatogram attached.
- Sequence assignment with the fragmentation spectrum showing b-ion and y-ion coverage.
- Lot number and synthesis date for traceability.
- Acceptance criteria and the analytical method used to generate each result.
“Incomplete COAs lacking evidence are insufficient to verify peptide sequences reliably. The COA is the contract between supplier and researcher, and it must include data proof, not just summary claims.” — COA interpretation standard for research peptides
Third-party peptide testing adds an independent layer of verification that internal supplier testing cannot provide. When a laboratory outside the manufacturing chain confirms the sequence and purity, the result carries greater credibility for publication and regulatory review.
How is peptide sequence verification applied in research and development?
Sequence verification is not a single-point check. It applies across the full lifecycle of a peptide, from initial synthesis through batch release and into active research use.
Synthetic peptide validation
For synthetic peptides under 50 residues, MS/MS is the standard confirmation method. A researcher ordering a custom peptide for a receptor binding study needs to confirm that the delivered compound matches the ordered sequence before running any assay. A single amino acid substitution at a critical binding residue can produce false negatives or false positives that corrupt an entire data set.
Therapeutic protein characterization
Peptide mapping is an established standard for batch release and product characterization of therapeutic proteins, including monoclonal antibodies. Regulatory agencies require sequence confirmation as part of the biologic license application process. Peptide mapping data generated during development must be reproducible across batches, making the analytical method itself subject to validation.
Batch-to-batch consistency
Sequence verification supports batch-to-batch consistency by detecting synthesis errors before a new lot enters active use. The table below summarizes common research applications and the verification approach used in each.
| Research application | Primary verification method | Key quality metric |
|---|---|---|
| Synthetic peptide for assay development | MS/MS fragmentation | Full b/y-ion series coverage |
| Therapeutic protein batch release | Bottom-up peptide mapping | Sequence coverage greater than 95% |
| Cosmetic peptide formulation | MS/MS plus HPLC purity | Purity greater than or equal to 99% |
| Novel peptide characterization | De novo sequencing plus MS/MS | Confidence score and ion coverage |
Researchers working with research-grade peptides should request the full analytical package, not just the COA summary page, before committing a new lot to active experiments. The raw spectra tell a story that summary tables cannot.
Pro Tip: For cosmetic peptide applications, verify that the COA includes both sequence confirmation and HPLC purity data. Cosmetic-grade claims without MS/MS evidence do not meet the documentation standard for peer-reviewed research.
Key takeaways
Peptide sequence verification requires MS/MS fragmentation data, a complete Certificate of Analysis, and manual expert review to produce results that are reliable for research and regulatory purposes.
| Point | Details |
|---|---|
| MS/MS is the standard method | Tandem mass spectrometry provides definitive sequence confirmation for peptides under 50 residues. |
| COA must include raw data | A valid COA contains MS spectra and chromatograms, not just summary purity claims. |
| Retention time is not enough | Deletion variants can co-elute; only MS/MS fragmentation confirms true peptide identity. |
| Manual review prevents errors | Isobaric dipeptides and artifacts require expert inspection beyond automated software outputs. |
| Regulatory frameworks apply | ICH Q6A defines identity testing standards that MS/MS satisfies and HPLC retention time does not. |
The part of sequence verification that most researchers underestimate
After years of working with peptide data, the pattern I see most often is not a failure of instrumentation. It is a failure of review. Researchers receive a COA, see a purity figure above 99%, and move forward without examining the MS/MS spectrum. That decision is understandable under time pressure, but it is where errors enter the system.
The isobaric problem is real and underappreciated. Two dipeptide combinations can share identical masses and fool even well-configured software. I have seen sequence assignments accepted from automated reports that, on manual inspection, showed clear gaps in the ion series. Those gaps are not noise. They are the instrument telling you something is wrong.
My practical recommendation is to treat the fragmentation spectrum as the primary document and the purity figure as supporting context, not the other way around. If your supplier cannot provide the raw MS/MS spectrum, that is the answer you need. A complete peptide lab protocol should make spectral review a mandatory step before any new lot enters active use.
The researchers who catch sequence errors early are not the ones with the best instruments. They are the ones who read the data.
— Admin
Verified peptides and full documentation at Peptilab
Researchers who need sequence-confirmed peptides backed by complete analytical documentation will find Peptilab’s catalog built for exactly that standard.
Every peptide in the Peptilab catalog ships with a Certificate of Analysis that includes MS/MS sequence confirmation, HPLC purity data, and lot-specific traceability. Purity is guaranteed at greater than or equal to 99%, and all testing is conducted through third-party laboratories. Canadian fulfillment means no import delays and no gaps in the documentation chain. Whether your work involves metabolic research, therapeutic peptide characterization, or cosmetic formulation, Peptilab provides the analytical evidence your research requires. Review the peptide research catalog to find sequence-verified compounds matched to your application.
FAQ
What is peptide sequence verification?
Peptide sequence verification is the analytical process of confirming that a peptide’s amino acids are arranged in the correct order, typically using tandem mass spectrometry to generate a sequence-specific fragmentation pattern.
Why is MS/MS the preferred method for verifying peptide sequences?
MS/MS produces b-ion and y-ion series that map directly to the amino acid sequence, providing unambiguous identity confirmation that chromatographic methods alone cannot deliver.
What should a Certificate of Analysis include for sequence verification?
A complete COA must include MS/MS fragmentation data, an HPLC chromatogram with purity percentage, sequence assignment, lot number, and the acceptance criteria used for each test.
Can retention time confirm peptide identity?
Retention time cannot confirm peptide identity on its own. Deletion variants with similar hydrophobicity can co-elute, making MS/MS fragmentation the only reliable method for definitive confirmation.
What are isobaric peptides and why do they matter?
Isobaric peptides share identical or near-identical masses, such as the dipeptide pairs SA and GT, which can cause automated software to generate incorrect sequence assignments. Manual review of the MS/MS spectrum is required to resolve these cases accurately.