

Peptide stability testing methods are defined as the systematic analytical procedures used to assess how well a peptide maintains its structural integrity and biological activity under specified conditions. These methods encompass incubation protocols, precipitation techniques, and detection technologies such as RP-HPLC, LC-MS/MS, and size-exclusion chromatography. The stability analysis of peptides is critical for determining shelf life, supporting regulatory submissions, and confirming that research-grade materials meet purity thresholds before use. Common degradation pathways include oxidation of methionine and cysteine, deamidation of asparagine and glutamine, and truncation at the peptide termini. Catching these changes early protects both data quality and downstream research outcomes.
What are the main steps in peptide stability testing methods?
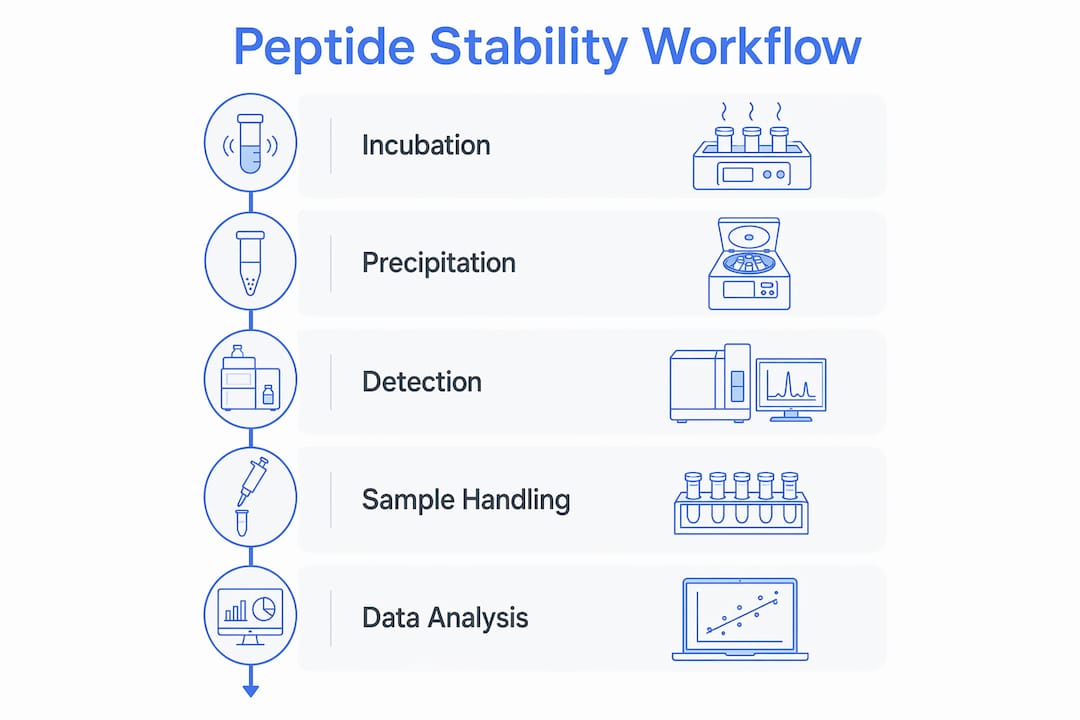
Peptide stability assays follow a three-phase workflow: incubation, precipitation, and detection. Each phase introduces variables that directly affect assay sensitivity and reproducibility. Choosing the wrong conditions in any phase can produce misleading stability profiles.
Phase 1: Incubation
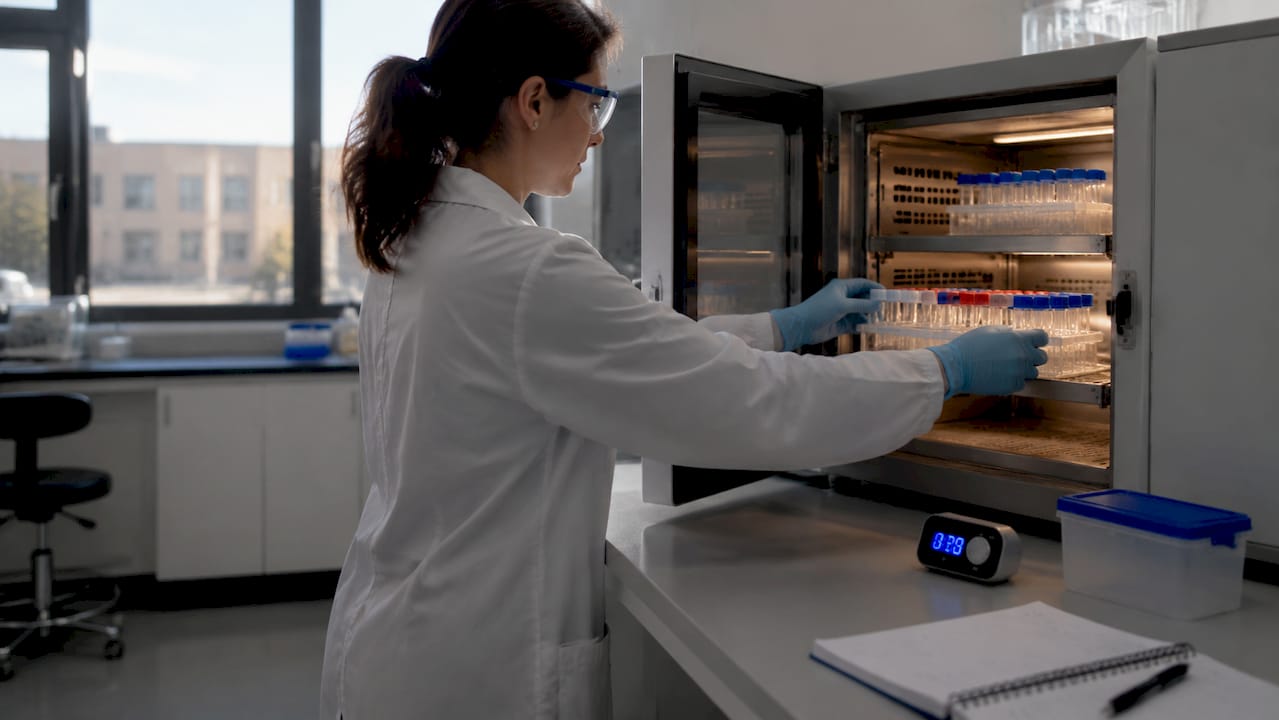
Incubation exposes the peptide to controlled stress conditions that simulate real-world degradation. Key variables include temperature, pH, solvent composition, and duration. Thermal stress studies typically run at 40°C, 60°C, or higher. Acidic and alkaline pH conditions probe hydrolytic susceptibility. Oxidative stress uses hydrogen peroxide or metal catalysts to accelerate methionine and cysteine oxidation.
Phase 2: Precipitation
Precipitation removes biological matrix components and concentrates the peptide for analysis. Acetonitrile and ethanol are the preferred precipitation solvents because they minimize peptide loss compared to trichloroacetic acid. Trichloroacetic acid can cause significant sample loss, which skews recovery calculations and underestimates degradation. Organic solvents preserve peptide integrity during cleanup and are the standard choice for plasma-based stability assays.
Phase 3: Detection
Detection identifies and quantifies intact peptide and degradation products. The most common detection sequence runs as follows:
- Inject the clarified sample onto a chromatographic column (RP-HPLC or SEC).
- Separate peptide species by hydrophobicity, size, or charge.
- Detect eluting peaks by UV absorbance at 214 nm or 280 nm.
- Couple the column to a mass spectrometer for structural confirmation.
- Integrate peak areas and calculate percent recovery relative to a reference standard.
Sample handling between phases matters as much as the phases themselves. Keeping samples on ice during preparation and minimizing freeze-thaw cycles prevents artifactual degradation before the assay even begins.
How do chromatographic techniques compare for peptide stability evaluation?
No single chromatographic method captures every degradation product a peptide can produce. Orthogonal methods with different separation principles are necessary to achieve comprehensive impurity coverage. The table below compares the three most widely used approaches.
| Method | Separation principle | Best for | Key limitation |
|---|---|---|---|
| RP-HPLC | Hydrophobicity | Purity profiling, oxidation, truncation | Poor resolution of charge variants |
| Size-exclusion chromatography (SEC) | Molecular size | Aggregation, oligomer detection | Low resolution for small impurities |
| Capillary electrophoresis (CE/icIEF) | Charge | Deamidation, isoform separation | Lower throughput than HPLC |
Reversed-phase HPLC is the workhorse of peptide stability evaluation. It separates peptides by hydrophobicity, making it highly sensitive to oxidation and truncation products that shift retention time. RP-HPLC is well-characterized, easy to validate, and accepted by regulatory agencies including the FDA and EMA.
Size-exclusion chromatography fills the gap RP-HPLC leaves open. SEC separates molecules by size, making it the primary tool for detecting aggregates and oligomers. Aggregation is a critical stability concern for therapeutic peptides, and SEC quantifies it directly without the denaturing conditions that RP-HPLC requires.
Capillary electrophoresis, including imaged capillary isoelectric focusing (icIEF), resolves charge variants that neither RP-HPLC nor SEC can separate. Deamidation of asparagine converts a neutral residue to a negatively charged aspartate, shifting the isoelectric point. icIEF detects that shift with high resolution.
Pro Tip: Run RP-HPLC and SEC in parallel on every forced degradation sample. The two methods together catch oxidation, truncation, and aggregation in a single experimental set, cutting the number of follow-up experiments needed.
What advanced detection methods enhance peptide degradation analysis?
Mass spectrometry is the definitive tool for peptide degradation analysis. LC-MS/MS and high-resolution mass spectrometry identify specific amino acid modifications and degradation fragments with structural precision that UV detection alone cannot provide. Coupling a liquid chromatography system to a mass spectrometer transforms a retention time shift into a confirmed molecular identity.
The core degradation pathways that mass spectrometry targets include:
- Oxidation: +16 Da mass shift on methionine or cysteine residues, detectable by extracted ion chromatography.
- Deamidation: +1 Da mass shift on asparagine or glutamine, converting them to aspartate or glutamate.
- Truncation: Loss of N-terminal or C-terminal residues, identified by fragment ion series in MS/MS spectra.
- Dimerization and cross-linking: Higher molecular weight species confirmed by intact mass analysis.
- Isomerization: Conversion of aspartate to isoaspartate, detectable by tandem mass spectrometry with characteristic mass shifts.
Peptide mapping integrates LC-MS/MS into a systematic workflow. The intact peptide is digested with a protease such as trypsin, and the resulting fragments are analyzed individually. Each fragment maps to a specific sequence position, so a mass shift on any fragment localizes the modification to a single amino acid. This level of resolution is required for regulatory submissions and forced degradation studies.
High-resolution mass spectrometry platforms, including Orbitrap and time-of-flight instruments, add another layer of confidence. They resolve isobaric species that unit-resolution instruments cannot distinguish. For GLP-1 receptor agonist peptides and other complex therapeutic sequences, high-resolution data is the standard for impurity profiling.
Pro Tip: Always run a blank gradient before your first sample injection in an LC-MS run. Background plasticizer peaks from tubing and fittings can overlap with low-abundance degradation products and produce false positives in impurity reports.
How to develop and validate stability-indicating methods for peptides
A stability-indicating method must resolve the main peptide peak from every degradation product the molecule can produce. Forced degradation studies under thermal, pH, and oxidative stress conditions generate the degradation products needed to challenge the method before validation begins.
The development sequence follows these steps:
- Define degradation pathways. Review the peptide sequence for susceptible residues: methionine, cysteine, asparagine, glutamine, and terminal residues.
- Design stress conditions. Apply thermal stress (40°C and 60°C for 1–4 weeks), acid and base hydrolysis (0.1 M HCl and NaOH), and oxidative stress (0.3% hydrogen peroxide for 24–48 hours).
- Select the primary method. Choose RP-HPLC as the baseline separation. Confirm that stressed samples produce new peaks that are resolved from the main peak.
- Add orthogonal methods. Incorporate SEC for aggregation and CE for charge variants. Each method must independently confirm the degradation products identified by RP-HPLC.
- Validate the method. Demonstrate specificity, linearity, accuracy, precision, and detection limits according to ICH Q2(R1) guidelines.
Validation parameters that regulators scrutinize most closely include:
- Specificity: The method must show that degradation product peaks do not co-elute with the main peak under any stress condition.
- Sensitivity: The limit of detection must be low enough to catch degradation products at the 0.1% level or below for drug-grade peptides.
- Precision: Repeatability and intermediate precision must meet predefined acceptance criteria across multiple analysts and instruments.
- Accuracy: Spiked degradation products must be recovered within the validated range.
A common pitfall is declaring a method stability-indicating based on one stress condition alone. Oxidative stress may produce a well-resolved impurity peak, but acid hydrolysis products may co-elute with the main peak on the same column. Testing all relevant stress conditions before validation prevents this failure mode.
What are best practices for reliable peptide stability testing?
Reliable peptide stability testing depends as much on sample handling as on instrument performance. Moisture contamination reduces long-term stability, while dry peptides remain stable at room temperature for days to weeks. Wet peptides degrade rapidly even at refrigerated temperatures.
Key practices that protect data integrity throughout the testing process:
- Store samples at the correct temperature. Cold storage at -20°C suppresses acid-driven hydrolysis and general degradation. Reserve room-temperature storage only for short-term working solutions.
- Minimize freeze-thaw cycles. Each cycle introduces mechanical stress and concentration gradients that accelerate aggregation. Aliquot samples before freezing.
- Use low-bind plasticware. Standard polypropylene tubes adsorb hydrophobic peptides. Low-bind tubes from suppliers such as Eppendorf or Thermo Fisher Scientific reduce surface loss significantly.
- Control solvent purity. HPLC-grade acetonitrile and water with less than 0.1% trifluoroacetic acid are the standard mobile phase components. Solvent impurities appear as ghost peaks that obscure low-level degradation products.
- Document every deviation. Any change in incubation time, temperature, or solvent lot number must be recorded. Undocumented deviations make out-of-specification results impossible to investigate.
Synthetic modifications also extend peptide stability in formulation. PEGylation, cyclization, and incorporation of D-amino acids each reduce susceptibility to proteolytic degradation. These modifications are worth considering early in the research cycle, not as a last resort after stability failures.
Key takeaways
Comprehensive peptide stability evaluation requires a combination of chromatographic separation, mass spectrometric detection, and rigorous sample handling, because no single method detects every degradation pathway a peptide can undergo.
| Point | Details |
|---|---|
| Three-phase assay workflow | Incubation, precipitation, and detection each introduce variables that affect reproducibility and sensitivity. |
| Orthogonal chromatography | RP-HPLC, SEC, and capillary electrophoresis together cover oxidation, aggregation, and charge variants. |
| Mass spectrometry for identification | LC-MS/MS localizes modifications to individual residues and confirms degradation product identity. |
| Forced degradation first | Stress studies under thermal, pH, and oxidative conditions must precede method validation. |
| Sample handling is critical | Cold storage, low-bind plasticware, and minimal freeze-thaw cycles prevent artifactual degradation. |
What I have learned from years of peptide stability work
The most consistent mistake I see in peptide stability workflows is treating RP-HPLC as a complete answer. Researchers run a single chromatographic method, see a clean profile, and conclude the peptide is stable. Then a mass spectrometry run reveals deamidation at asparagine-17 that the HPLC column simply could not resolve from the main peak. The peptide was degrading the entire time. The method just was not sensitive enough to show it.
The second lesson is about timing. Forced degradation studies feel like extra work at the start of a project. They are not. Running thermal and oxidative stress experiments in the first two weeks of method development saves months of troubleshooting during validation. Every degradation product you identify early becomes a known quantity you can monitor and control.
Emerging detection technologies are shifting what is possible. Ion mobility mass spectrometry now separates conformational isomers that share identical mass and chromatographic retention. For cyclic peptides and constrained scaffolds, this matters enormously. Researchers working on research peptides in Canada and elsewhere are beginning to incorporate ion mobility data into stability packages for regulatory filings. The field is moving faster than most standard operating procedures reflect.
The practical takeaway is this: build your stability testing strategy around the peptide’s specific sequence vulnerabilities, not around the instruments you already own. Let the chemistry drive the method selection.
— Admin
Quality peptides and testing support from Peptilab
Stability testing is only as reliable as the peptide you start with. Peptilab supplies research-grade peptides verified at greater than 99% purity through third-party analytical testing, with full certificates of analysis available for every product. Each COA documents purity data, mass confirmation, and lot-specific analytical results, giving your stability studies a documented baseline from day one.
Peptilab ships from Canada with no import delays, making it a practical source for laboratories that need consistent, well-documented peptide material for long-term stability testing programs. Whether your work involves metabolic research, anti-aging formulations, or cosmetic peptide development, Peptilab’s full peptide catalog covers the sequences most commonly used in stability and performance studies.
FAQ
What does a peptide stability assay measure?
A peptide stability assay measures the rate and extent of chemical and physical degradation under defined conditions. It quantifies intact peptide recovery and identifies degradation products such as oxidized, deamidated, or truncated species.
Which chromatographic method is best for peptide purity testing?
RP-HPLC is the primary method for purity profiling because it separates peptides by hydrophobicity and detects oxidation and truncation products reliably. SEC and capillary electrophoresis are added as orthogonal methods to cover aggregation and charge variants.
How does LC-MS/MS improve degradation analysis?
LC-MS/MS identifies the exact amino acid residue affected by degradation by measuring mass shifts on individual peptide fragments. This structural specificity is not achievable with UV detection alone.
Why are forced degradation studies required before method validation?
Forced degradation studies generate the actual degradation products the method must resolve. Without them, specificity cannot be demonstrated, and the method cannot be confirmed as stability-indicating under ICH Q2(R1) guidelines.
How should peptides be stored to minimize degradation before testing?
Dry peptides should be stored at -20°C in sealed, moisture-free containers. Cold storage suppresses hydrolysis and general degradation, while moisture contamination accelerates breakdown even at refrigerated temperatures.