

Peptide lead optimisation is the iterative, multi-parameter process of converting a confirmed hit into a research-ready lead compound by simultaneously improving potency, metabolic stability, selectivity, and pharmacokinetic (PK) exposure. It is not a single experiment. It is a structured feedback cycle where each round of data narrows the chemical space and informs the next design decision. Three outputs define a completed optimisation round:
- A fully characterised lead sequence with defined binding affinity and cellular activity
- A certificate of analysis (COA) confirming purity, ideally >99% by HPLC and MS
- A documented go/no-go decision against pre-defined PK and potency thresholds
Table of Contents
- Why does lead optimisation matter for cosmetic and biomedical research?
- What does the tiered optimisation workflow look like?
- Which SAR strategies work best for peptides?
- What metrics should you track across optimisation rounds?
- How should you use AI and cheminformatics tools?
- Practical sourcing and QA considerations for Canadian labs
- How do you build a go/no-go decision framework?
- A short worked example: from hit to lead in two rounds
- Key takeaways
- The part most researchers get wrong
- Research-grade peptides for Canadian labs: Peptilab
- Selected further reading
Why does lead optimisation matter for cosmetic and biomedical research?
Skipping or shortcutting optimisation has direct downstream consequences. Poorly characterised leads produce irreproducible assay data, undermine translational confidence, and create formulation problems that surface late when they are expensive to fix. For cosmetic researchers, a peptide with a 20-minute serum half-life will not survive a stability challenge in an aqueous formulation, regardless of how impressive its receptor-binding data looks; products like Perimenopause premium collagen peptides highlight the importance of optimised peptide stability in topical formulations. For biomedical teams, a lead that lacks selectivity will confound mechanistic conclusions and complicate any ethics review.
The scientific goals connect directly to project outcomes. Target engagement requires adequate binding kinetics. Translational relevance requires a half-life matched to the assay window. Permeability determines whether a peptide reaches an intracellular target or stays confined to the extracellular compartment. Cosmetic peptide efficacy depends on all three, as detailed in cosmetic efficacy testing protocols.
Pro Tip: Run stability assays in the first round, not the last. A peptide that degrades in two minutes in serum will waste every subsequent chemistry cycle spent improving its potency.
What does the tiered optimisation workflow look like?
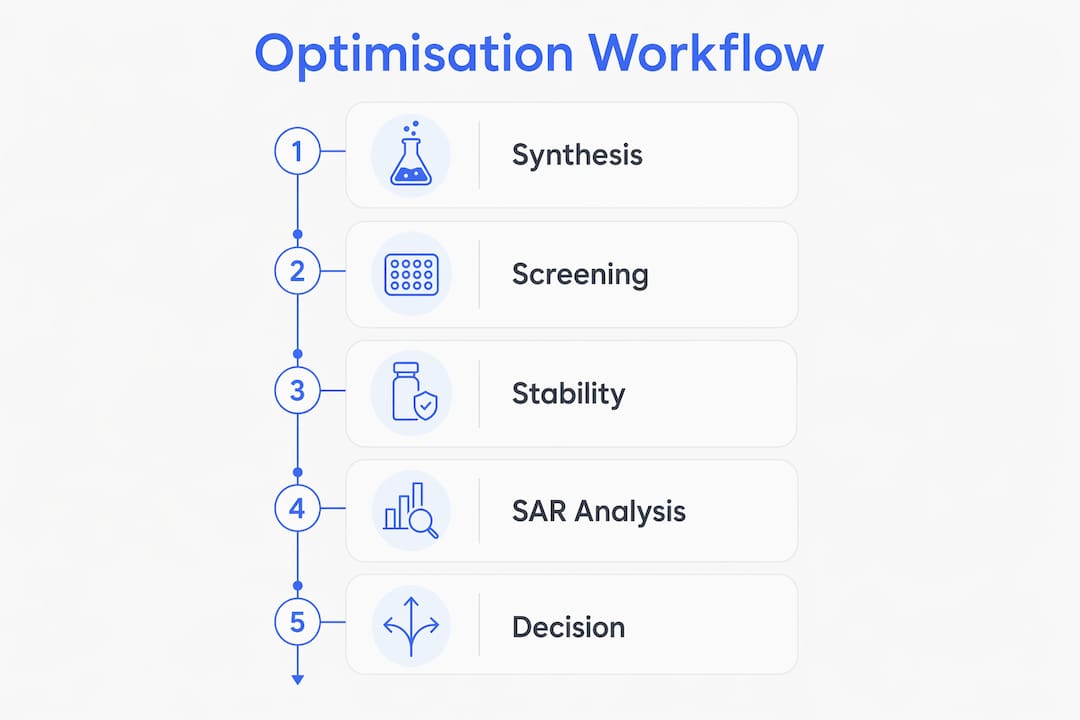
The standard pipeline moves from low-cost biophysical assays toward higher-cost mechanistic and in vivo work, using failure at each tier to redirect chemistry before resources are committed.
- Tier 1 — biophysical binding: Surface plasmon resonance (SPR), biolayer interferometry (BLI), and isothermal titration calorimetry (ITC) measure binding kinetics (kon, koff) and equilibrium affinity (KD). Kinetics matter because a fast off-rate (high koff) predicts short target residence time, which often correlates with reduced in vivo efficacy even when KD looks acceptable.
- Tier 2 — cellular activity and permeability: EC50/IC50 in relevant cell lines, Caco-2 or PAMPA permeability (Papp), and efflux ratio profiling. A peptide that fails permeability here redirects chemistry toward cyclisation or backbone modification before Tier 3 is attempted.
- Tier 3 — mechanistic and PK validation: Cellular thermal shift assay (CETSA) for target engagement, ex vivo or simple in vivo PK models for half-life and clearance. This tier generates the go/no-go data that determines whether a lead advances. A tiered evaluation pipeline aligns biological half-life targets with physiological clearance rates.
| Assay | Primary metric | Typical throughput | Decision threshold |
|---|---|---|---|
| SPR / BLI | KD, kon, koff | Medium (microplate) | KD ≤ target-specific nM range |
| ITC | ΔH, KD | Low (1–4/day) | Confirms SPR; flags enthalpy/entropy |
| Caco-2 / PAMPA | Papp (cm/s) | High (plate-based) | Papp > 1×10⁻⁶ cm/s (passive) |
| Cell potency | EC50 / IC50 | Medium | ≤ 10× Tier 1 KD |
| Serum stability | t½ (min/h) | Medium | Project-defined; typically >2 h |
| CETSA | Tm shift (°C) | Low-medium | ≥ 2°C shift confirms engagement |
A focused optimisation campaign of 10–20 analogues typically runs two to four weeks per cycle from synthesis receipt to Tier 2 data.
Which SAR strategies work best for peptides?
Alanine scanning and matched molecular pair (MMP) analysis are the most effective starting points. Alanine scans identify critical contact residues (substitution cost >2 kcal mol⁻¹ flags a pharmacophoric position); permissive positions become handles for ADME-improving modifications. MMP networks then let you compare pairs of analogues that differ by a single defined change, keeping SAR interpretable.
Common modifications and what they primarily address:
- Cyclisation / macrocycles: conformational constraint improves selectivity and protease resistance, but larger macrocycles often trade permeability for potency
- Backbone N-methylation: blocks amide bond proteolysis and reduces hydrogen-bond donors, improving membrane permeability
- D-amino acid substitution: disrupts protease recognition without large conformational cost; extends plasma half-life from minutes to hours
- Stapling (hydrocarbon or lactam): locks helical conformation, improves cell penetration for helical peptides
- Lipidation / fatty acid conjugation: extends half-life via albumin binding (semaglutide is the canonical example)
- PEGylation: reduces renal clearance and immunogenicity; can reduce receptor affinity at high MW
Pro Tip: Design your library so each analogue differs from the parent by exactly one modification. Clean MMP analysis becomes impossible when two variables change simultaneously, and you lose the ability to assign cause to effect.
Multiparameter optimisation (MPO) is the governing concept here. A potency gain that costs three log units of solubility is rarely a net improvement. Every modification decision should be scored across all parameters simultaneously, not sequentially.
What metrics should you track across optimisation rounds?
The most critical quantitative readouts are binding kinetics (KD, kon, koff), cellular potency (EC50/IC50), metabolic stability (serum t½), passive permeability (Papp), and selectivity scores against off-target panels.
- KD (nM): equilibrium dissociation constant from SPR/ITC; improvement = lower value toward target range
- EC50/IC50 (nM–µM): functional potency in cell line; improvement = shift toward lower concentration
- Serum t½ (min or h): half-life in human or species-matched serum; improvement = longer duration
- Papp (cm/s): passive permeability from Caco-2 or PAMPA; improvement = higher value for intracellular targets
- Selectivity ratio: ratio of off-target IC50 to on-target IC50; improvement = ratio >100× preferred
For peptide stability testing, pairing serum incubation with MS fragmentation identifies which bonds are cleaved, directing subsequent chemistry.
| Metric | Assay | Typical decision benchmark |
|---|---|---|
| KD | SPR / ITC | ≤ 100 nM for most therapeutic targets |
| EC50 | Cell-based reporter or functional | ≤ 1 µM to advance to Tier 3 |
| Serum t½ | Serum stability + LC-MS | > 2 h (project-specific) |
| Papp | Caco-2 A→B | > 1×10⁻⁶ cm/s for oral/topical targets |
| Selectivity | Counter-screen panel | ≥ 100-fold over nearest off-target |
How should you use AI and cheminformatics tools?
Use them to prioritise and reduce synthesis burden, not to replace wet-lab validation. AI tools reduce candidate numbers to test but function best when used to propose substitutions that are then confirmed by orthogonal biophysical and cellular assays.
Practical in silico tactics:
- AlphaFold3 / RFdiffusion: structure prediction and cyclic peptide design to generate binding hypotheses
- Free energy perturbation (FEP): pre-screen single-point mutations for relative affinity before synthesis
- MMP analysis with PepFuNN: open-source toolkit combining protein and small-molecule tools for peptide alignments and non-canonical amino acid handling
- PepSeA: sequence alignment and SAR visualisation for cyclic and cross-linked peptides with non-natural residues
- mPARCE: iterative computational protocol for optimising modified peptides via single-point mutations scored by consensus docking functions
Closed-loop predict–synthesise–test–retrain workflows compress timelines from years to months when synthesis, screening, and structural analysis are tightly integrated. The key word is “tightly”: a compute prediction that sits in a queue for six weeks delivers no timeline benefit.
Pro Tip: Prioritise in silico substitutions that are low-uncertainty and testable in your existing assay panel. A prediction that requires a new assay to verify is a bottleneck, not a shortcut.
Practical sourcing and QA considerations for Canadian labs
Canadian researchers should prioritise suppliers that provide COAs with explicit analytical methods, third-party purity verification (>99% by HPLC and MS where required), domestic fulfilment, and clear storage and handling instructions. Import delays for peptides crossing the US–Canada border can add one to three weeks to a procurement cycle, which disrupts iterative optimisation cadences.
Purchasing and QA checklist:
- COA with HPLC chromatogram and MS confirmation of molecular weight
- Stated analytical method (column, gradient, detection wavelength)
- Endotoxin status (LAL or equivalent) for cell-based or in vivo work
- Batch size options matched to your assay scale (1 mg to 100 mg typical for optimisation)
- Cold-chain shipping confirmation and recommended storage temperature
- Lead time from order to delivery (domestic Canadian suppliers: typically 3–7 business days)
Cost drivers for small optimisation runs include scale of synthesis, incorporation of non-canonical residues (which require custom Fmoc building blocks), cyclisation chemistry, and analytical QC scope. A focused 10-analogue panel at 1–5 mg scale is a realistic starting point for Tier 1–2 screening.
Pro Tip: Order your next synthesis round before your current assay data is fully analysed. The two-to-four-week synthesis lead time and the one-to-two-week assay turnaround can run in parallel if procurement is planned a cycle ahead.
How do you build a go/no-go decision framework?
Make go/no-go decisions using a pre-defined, weighted multi-parameter score across potency, stability, permeability, and manufacturability. Setting thresholds before data arrives removes confirmation bias from the decision.
Steps to build a scoring rubric:
- Define minimum acceptable thresholds for each parameter (e.g., EC50 ≤ 1 µM, serum t½ > 2 h, Papp > 1×10⁻⁶ cm/s)
- Assign relative weights based on project priority (stability-first for cosmetic leads; potency-first for receptor-targeted biomedical leads)
- Score each analogue numerically; any compound failing a hard minimum is eliminated regardless of total score
- Rank passing compounds by weighted total; advance the top two to three into the next cycle
- When PK data shows adequate exposure, priorities further potency optimisation and shift resources to manufacturability and formulation compatibility
A compound that passes all hard minimums but ranks mid-table on the weighted score is a better investment than a potency champion with a fatal stability flaw.
A short worked example: from hit to lead in two rounds
Starting hit: KD = 850 nM (SPR), EC50 = 4.2 µM (cell reporter), serum t½ = 8 minutes. Primary problem: proteolytic instability.
- Round 1 modification: MS cleavage mapping identified a Lys-Pro bond as the primary protease site. Substituted D-Lys at that position and added backbone N-methylation at the adjacent residue. Synthesised six analogues.
- Round 1 results: Best analogue: KD = 310 nM, EC50 = 1.1 µM, serum t½ = 2.4 h. Permeability unchanged (Papp = 0.8×10⁻⁶ cm/s). Decision: advance, address permeability in Round 2.
- Round 2 modifications: Introduced a lactam cyclisation between positions 3 and 7 to constrain conformation and reduce hydrogen-bond donors. Four analogues synthesised.
- Round 2 results: Best cyclic analogue: KD = 180 nM, EC50 = 0.6 µM, serum t½ = 3.1 h, Papp = 1.4×10⁻⁶ cm/s. All hard minimums met. Go decision issued.
Each round from synthesis receipt to Tier 2 data took approximately three weeks, putting the full two-round campaign at roughly six weeks.
Key takeaways
Peptide lead optimisation requires a tiered, multi-parameter workflow where experimental data from each round drives the next design decision, and no single metric determines advancement.
| Point | Details |
|---|---|
| Optimise all parameters together | Potency, stability, permeability, and selectivity must be tracked simultaneously, not sequentially. |
| Run the tiered pipeline in order | Tier 1 biophysical data gates Tier 2 cell assays; Tier 3 PK is reserved for compounds that pass both. |
| Use AI to narrow, not decide | Computational tools like AlphaFold3 and PepFuNN reduce synthesis burden but require wet-lab confirmation. |
| Set go/no-go thresholds before data arrives | Pre-defined weighted scores remove confirmation bias and keep campaigns on schedule. |
| Peptilab for Canadian sourcing | Peptilab supplies research-grade peptides with COAs, >99% purity verification, and domestic Canadian fulfilment. |
The part most researchers get wrong
The most persistent mistake in peptide lead optimisation is treating it as a potency problem. Teams spend three or four rounds chasing a lower EC50 while ignoring a serum half-life that will disqualify the compound the moment it enters any PK model. The multidimensional balance between potency, stability, and bioavailability is the actual challenge, and a moderately potent, stable, bioavailable candidate consistently outperforms a potency champion with fatal ADME liabilities.
The shortcut that saves the most time is early MS mapping of protease cleavage sites. Routine MS mapping of cleavage fragments lets chemists place D-amino acids or N-methylation at vulnerable positions before committing to in vivo work. Doing this in Round 1 rather than Round 3 typically saves four to six weeks of wasted synthesis.
Research-grade peptides for Canadian labs: Peptilab
Canadian research teams running optimisation campaigns need a supplier that keeps pace with iterative cycles, not one that adds three weeks of import uncertainty to every round. Peptilab supplies research-grade peptides with COAs, HPLC and MS purity confirmation (>99% where specified), and domestic Canadian fulfilment, which means your next synthesis round arrives on a predictable schedule.
The catalogue covers peptides for metabolic research, cellular studies, cosmetic applications, and recovery formulations, alongside laboratory essentials including bacteriostatic water, syringes, and alcohol wipes. When placing an order, request the analytical method used in the COA, endotoxin status for any cell-based or in vivo work, and confirmed cold-chain handling. Peptilab also carries formulation supplies that support moving leads toward preclinical testing.
Disclosure: this article is published by Peptilab. Verify COAs and batch testing documentation before purchase, regardless of supplier.
Browse the full catalogue at peptilab.ca/product-category/all-peptides to find the peptide grade and batch size that fits your current optimisation round.
Selected further reading
- Protocol for iterative optimisation of modified peptides bound to protein targets (mPARCE) — detailed computational workflow for single-point mutation optimisation using Rosetta; cite in methods when using structure-guided design.
- From lead to market: chemical approaches to transform peptides into therapeutics — review covering PK optimisation, medicinal chemistry modifications, and case studies including semaglutide and MK-0616.
- Peptides as programmable molecular scaffolds (RSC Chemical Biology) — comprehensive review of modifications, AI-guided design, and closed-loop workflows.
- Lead discovery and optimisation strategies for peptide macrocycles — focused treatment of cyclisation strategies, permeability trade-offs, and macrocycle SAR.
- PepFuNN: open-source toolkit for peptide in silico analysis — practical software for MMP analysis, sequence alignment, and non-canonical amino acid handling.
- PepSeA: peptide sequence alignment and visualisation for lead optimisation — open-source tool for SAR visualisation of cyclic and cross-linked peptides with non-natural residues.
- Structure-based molecular modelling in SAR analysis and lead optimisation (review) — balanced assessment of where in silico SAR methods add value and where wet-lab confirmation remains mandatory.
- Protease degradation mapping and peptide stability — practical guidance on MS-based cleavage site identification and corrective substitution strategies.