

Peptide mass spectrometry is an analytical technique that identifies and characterises peptides by ionising them, separating the resulting ions by their mass-to-charge ratio (m/z), and converting that measurement into an intact molecular weight or fragment pattern. The result: confirmed identity, detected post-translational modifications (PTMs), and an orthogonal check that HPLC purity data alone cannot provide. An HPLC area percent near 98% does not guarantee equivalent peptide purity by mass, because counterions, residual solvents, and salts can inflate apparent purity. Mass spectrometry closes that gap by measuring the molecule itself.
Two things to confirm on any COA before accepting a peptide batch:
- MS method and instrument (ESI-MS, MALDI-TOF, or LC-MS) with the observed mass, theoretical mass, and stated tolerance
- Third-party verification from an ISO/IEC 17025-accredited laboratory, with a batch-specific spectrum image and an explicit pass/fail decision
Peptilab supplies research-grade and cosmetic peptides in Canada with batch-specific COAs that include MS data, giving researchers and formulators a documented starting point.
Table of Contents
- How peptide mass spectrometry works
- Which MS method fits your peptide work?
- What to do before sending peptides for MS
- How to read your MS results
- Where peptide MS matters most for your work
- Known limits of peptide MS and how to fix common problems
- What a COA should contain and why accreditation matters
- In-house analysis versus contract labs in Canada
- Key takeaways
- A practical word on integrating MS into your QC workflow
- Peptilab: research-grade peptides with MS-backed documentation for Canadian labs
- Useful sources and further reading
How peptide mass spectrometry works
Every mass spectrometry experiment follows the same four-step sequence, regardless of instrument type.
1. Sample introduction. Peptides enter the instrument either directly, spotted onto a target plate (MALDI), or via an LC column coupled online to the ion source (LC-ESI).
2. Ionisation. The ion source converts neutral peptide molecules into gas-phase ions. Electrospray ionisation (ESI) sprays a liquid sample through a high-voltage capillary, producing multiply charged ions. MALDI fires a laser at a co-crystallised matrix, generating predominantly singly charged ions.
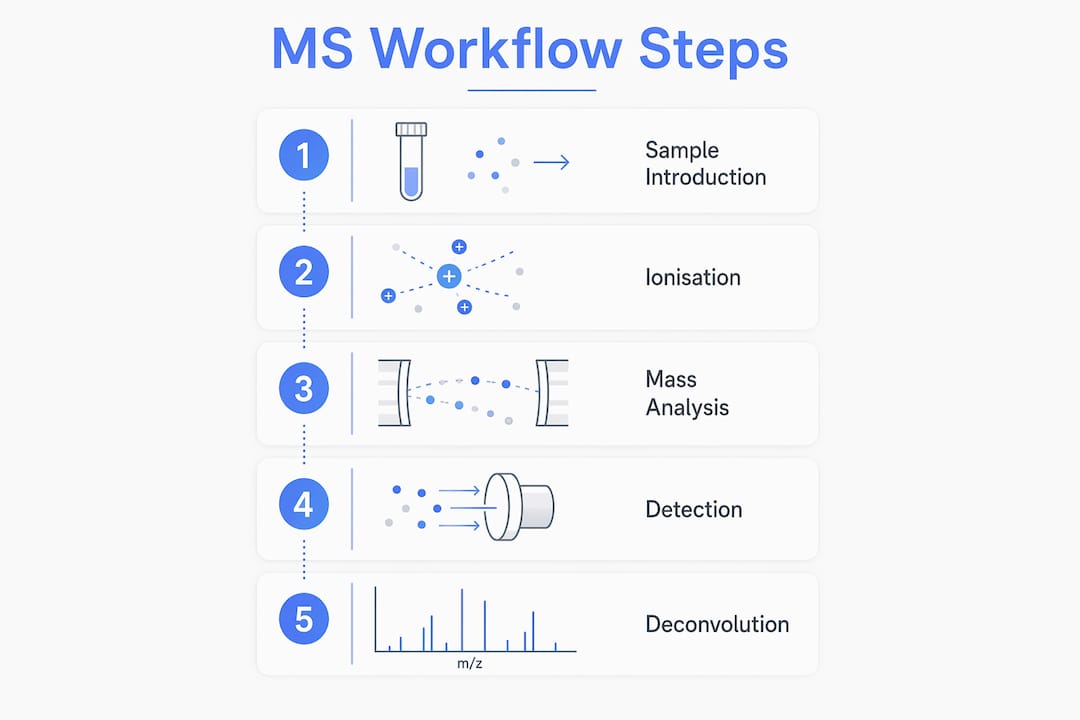
3. Mass analysis. A mass analyser separates ions by m/z. Quadrupoles filter ions sequentially; time-of-flight (TOF) analysers measure how long ions take to travel a fixed distance; Orbitraps measure the frequency of ion oscillation around a central electrode. Each approach trades speed, resolution, and cost differently.
4. Detection and deconvolution. The detector records ion abundance at each m/z value. For ESI data, deconvolution software calculates the neutral molecular weight from the pattern of multiply charged peaks. A coherent ESI envelope that resolves cleanly to one intact mass is a strong indicator of a single correct peptide in the sample.
Pro Tip: Before ESI analysis, desalt your peptide solution using a ZipTip or solid-phase extraction (SPE) cartridge. Even low millimolar salt concentrations suppress ionisation and broaden charge-state envelopes, making deconvolution unreliable. Aim for less than 10 mM ammonium acetate as your only buffer salt.
Which MS method fits your peptide work?
Choosing the right approach depends on your throughput needs, sample matrix, and whether you need sequence-level data or just an intact mass.
| Method | Ionisation | Charge behaviour | Best sample types | Throughput | Typical mass accuracy |
|---|---|---|---|---|---|
| MALDI-TOF | Laser/matrix | Singly charged [M+H]⁺ | Synthetic peptides, salts tolerated, <10 kDa | High | ±1 Da |
| ESI (direct infusion) | Electrospray | Multiply charged | Clean solutions, desalted | Medium | <0.1 Da (high-res) |
| LC-ESI-MS | Electrospray + LC | Multiply charged | Complex mixtures, impurity profiling | Medium | <0.1 Da or <5 ppm |
| LC-MS/MS (tandem) | Electrospray + LC | Multiply charged | PTM localisation, sequence confirmation | Lower | <5 ppm |
MALDI-TOF produces predominantly singly charged ions, making spectra straightforward to interpret: the major peak sits at approximately molecular weight + 1.008 Da. That simplicity, combined with tolerance for moderate salt levels, makes it the standard choice for rapid identity checks on COAs. LC-ESI-MS couples chromatography to MS and delivers both purity and identity in a single run, resolving co-eluting impurities that UV detection alone cannot distinguish. Typical acceptable tolerances are approximately ±1 Da for MALDI-TOF and a fraction of a Dalton (or a few ppm) for high-resolution Q-TOF or Orbitrap instruments. The instrument type must appear on the COA so the tolerance criterion is interpretable.
For cosmetic formulators, a quick MALDI or LC-ESI intact mass check on incoming batches covers most identity needs. Reserve LC-MS/MS for PTM localisation or when sequence ambiguity is suspected.
What to do before sending peptides for MS
Poor sample preparation is the leading cause of rejected submissions and uninterpretable spectra. Follow this checklist before shipping to a contract lab.
- Dissolve in a compatible solvent. Use water, acetonitrile/water mixtures, or dilute acetic acid. Avoid DMSO unless the lab confirms compatibility.
- Remove salts and detergents. Use a ZipTip C18 or SPE cartridge. Phosphate buffers and SDS suppress ionisation severely.
- Set the right concentration. Most labs target 0.1–1 mg/mL for intact peptides; confirm with the lab before submission.
- Include a reference standard or internal calibrant. A known-mass peptide run alongside your sample validates instrument calibration.
- Label clearly. Include peptide name, sequence, expected mass, solvent, batch number, and requested analyses on both the tube and the submission form.
- Attach chain-of-custody documentation. For Canadian contract labs, this means a sample receipt form, your COA request, and any acceptance criteria you need the lab to apply.
- Enzymatic digestion is rarely needed for synthetic peptides. Intact mass analysis is sufficient unless you are working with a complex protein mixture or need sequence coverage data.
Pro Tip: Trifluoroacetic acid (TFA) used in peptide synthesis and HPLC purification leaves residual TFA adducts (+114 Da) that can confuse mass assignments. If your peptide was purified by RP-HPLC with TFA, request a brief solvent exchange to 0.1% formic acid before ESI analysis, or note TFA content on your submission form so the analyst can account for it.
See Peptilab’s peptide lab protocol guide for a detailed sample handling reference.
How to read your MS results
The core comparison is simple: observed mass versus theoretical mass, within the instrument’s stated tolerance. If those agree, the peptide passes identity. If they do not, the magnitude and sign of the discrepancy tell you where to look.
Charge state calculation from adjacent ESI peaks follows: n = (m₂ − 1.008) / (m₁ − m₂), where m₁ and m₂ are adjacent m/z peaks. Modern software automates this, but understanding the formula helps when deconvolution produces unexpected results.
| Mass shift | Likely cause | Suggested follow-up |
|---|---|---|
| +16 Da | Methionine or tryptophan oxidation | Check storage conditions; repeat after fresh dissolution |
| +1 Da | Deamidation (Asn → Asp or Gln → Glu) | LC-MS/MS to localise site |
| +22 Da | Sodium adduct | Desalt and repeat |
| +38 Da | Potassium adduct | Desalt and repeat |
| −18 Da | Water loss (cyclisation) | LC-MS/MS; check synthesis report |
| — | Deletion or sequence error | Request LC-MS/MS and synthesis records |
Common mass shifts indicate likely causes: small shifts (+16, +22) point to adducts or oxidation; large discrepancies of tens to hundreds of daltons typically indicate a synthesis deletion or sequence error rather than a simple adduct. A COA report should list the instrument and method, theoretical mass, observed mass, tolerance, a spectrum image, and an explicit pass/fail statement.
Where peptide MS matters most for your work
Mass spectrometry supports several distinct jobs in research and cosmetic formulation workflows.
- Identity confirmation on incoming batches. MS is the gold-standard technique for confirming peptide identity; it can distinguish peptides differing by a single amino acid, something HPLC cannot do.
- Purity verification when coupled to LC. LC-MS assigns molecular weights to each chromatographic peak, so a co-eluting impurity with the same retention time as your target peptide is no longer invisible.
- PTM localisation for functional studies. MS/MS b/y ion series pinpoint modification sites on phosphorylated, glycosylated, or acetylated peptides critical to mechanism studies.
- Stability testing for cosmetic formulations. Comparing intact mass spectra of a peptide ingredient before and after accelerated stability conditions reveals degradation products that UV absorbance misses entirely.
- Batch consistency checks. Repeated intact mass confirmation across production lots catches synthesis drift before it reaches a formulation.
For cosmetic formulators exploring topical peptide applications, evidence-based peptide research increasingly relies on MS-confirmed identity as a prerequisite for efficacy claims. Peptilab’s peptide chromatography guide explains how LC and MS data complement each other in purity reporting.
Known limits of peptide MS and how to fix common problems
Ion suppression is the most common failure mode. Salts, detergents, and polymer contaminants ionise efficiently and crowd out peptide signal. The fix is always desalting first. If signal remains low after desalting, concentrate the sample and reduce the injection volume.
Incomplete fragmentation in MS/MS is the second common issue, particularly for peptides longer than 20 residues or those with many basic residues. Switching from collision-induced dissociation (CID) to electron transfer dissociation (ETD) often improves sequence coverage on larger or heavily modified peptides.
Ambiguous adducts (+22 Da sodium, +38 Da potassium) are resolved by desalting and repeating. If the shift persists after desalting, request a high-resolution repeat on a Q-TOF or Orbitrap to confirm the exact mass and rule out a real modification.
Pro Tip: For challenging peptides with poor fragmentation coverage, ask the lab to run both CID and ETD on the same precursor. The complementary fragment sets often fill gaps that either method alone leaves, particularly around proline residues, which block CID fragmentation at that bond.
What a COA should contain and why accreditation matters
A COA that lists only HPLC data leaves identity unverified. Insist on MS data before accepting a high-purity claim for research or formulation work. The MS section of a defensible COA must include:
- Analytical method (ESI-MS, MALDI-TOF, or LC-MS)
- Instrument model and resolution class
- Theoretical monoisotopic or average mass
- Observed mass with units (Da)
- Stated tolerance and whether the result passes or fails
- Spectrum image (raw or annotated)
- Lot/batch number, analyst name, and analysis date
ISO/IEC 17025 accreditation means the laboratory’s methods have been independently validated and its measurement uncertainty is documented. For Canadian contract labs, ask specifically whether their peptide MS method is within the accreditation scope, not just whether the lab holds a general accreditation certificate. Chain-of-custody documentation, from sample receipt through analysis to report issue, protects result integrity and is required for any regulatory or marketing claim. Peptilab provides batch-specific QC documentation with every order, giving you a documented baseline before you send samples for further testing.
In-house analysis versus contract labs in Canada
For most Canadian research groups and cosmetic formulators, contract analysis is the practical starting point. Here is how to evaluate your options.
- Confirm method experience. Ask whether the lab routinely runs synthetic peptides, not just tryptic digests from proteomics workflows. The sample prep and acceptance criteria differ.
- Verify accreditation scope. ISO/IEC 17025 accreditation must cover the specific method (MALDI-TOF or LC-MS/MS) you need, not just the lab’s general operations.
- Clarify turnaround and pricing. Intact mass by MALDI or LC-ESI typically turns around in 2–5 business days at Canadian contract labs; LC-MS/MS with PTM mapping takes longer. Pricing varies by method and sample count.
- Specify your acceptance criteria upfront. Send the peptide sequence, expected mass, solvent matrix, requested analyses, and the tolerance you will accept. Ambiguity at submission leads to reports you cannot use.
- Weigh in-house purchase carefully. A benchtop MALDI or LC-MS system costs tens of thousands to hundreds of thousands of dollars in capital, plus maintenance contracts and trained staff. For labs running fewer than several dozen samples per month, contract analysis is almost always more cost-effective.
For peptide sequence verification needs beyond intact mass, the same submission checklist applies but add a request for MS/MS and specify which PTMs to look for.
Key takeaways
Mass spectrometry confirms peptide identity by measuring molecular weight directly; HPLC measures chromatographic purity and cannot substitute for that identity check.
| Point | Details |
|---|---|
| MS vs HPLC | MS confirms molecular identity; HPLC measures chromatographic purity — you need both. |
| COA minimum fields | Method, instrument, theoretical mass, observed mass, tolerance, spectrum image, and pass/fail. |
| Tolerance by instrument | MALDI-TOF: ±1 Da; high-resolution Q-TOF or Orbitrap: <0.1 Da or <5 ppm. |
| When to request MS/MS | Request tandem MS when PTMs, sequence ambiguity, or large mass discrepancies are present. |
| Peptilab sourcing | Peptilab supplies Canadian researchers and formulators with batch-specific COAs including MS data and third-party verification. |
A practical word on integrating MS into your QC workflow
The gap between what researchers say they do and what they actually do with MS data is wider than most labs admit. Intact mass confirmation on every incoming batch is not excessive rigour; it is the minimum defensible standard when your downstream work depends on the peptide being what the label says. For cosmetic formulators, that standard matters even more because a stability claim or an efficacy argument built on a misidentified ingredient is a liability, not just a scientific error.
The cost argument against routine MS checks has weakened considerably. Contract labs in Canada now offer intact mass by LC-ESI for a fraction of what it cost a decade ago, and the turnaround fits most procurement cycles. Where I see formulators go wrong is reserving MS for troubleshooting rather than using it as a gate. By the time a formulation batch fails, you have already spent the time and materials. Run the check on receipt, not after the fact.
For PTM-sensitive work, the calculus is different. LC-MS/MS adds cost and time, and not every batch warrants it. A reasonable middle ground: intact mass on every lot, LC-MS/MS on the first lot from a new supplier or when the application depends on a specific modification being present or absent.
Peptilab: research-grade peptides with MS-backed documentation for Canadian labs
The practical bottleneck for most Canadian researchers and formulators is not understanding mass spectrometry — it is sourcing peptides that arrive with the documentation to support it.
Peptilab ships research-grade and cosmetic peptides domestically across Canada, with no import delays and batch-specific COAs that include MS identity data and third-party verification. Every order comes with the lot number, observed mass, and spectrum documentation you need to satisfy an ISO/IEC 17025-aligned QC checklist. For formulators, Peptilab’s cosmetic peptide sourcing guide walks through what to request from any supplier before committing to a batch. For researchers working on stability or efficacy studies, the peptide cosmetic stability workflow pairs directly with the MS-based degradation testing described above. Browse the full catalogue or contact Peptilab for batch documentation and sourcing guidance specific to your application.
Useful sources and further reading
“Mass spectrometry has displaced Edman degradation because it is much more sensitive and can fragment peptides in seconds instead of hours or days. MS does not require proteins or peptides to be purified to homogeneity and has no problem identifying blocked or otherwise modified proteins.” — The ABCs of Peptide Sequencing by MS, Winthrop University
Key references used in this article:
- The ABCs of Peptide Sequencing by MS — foundational explanation of MS workflow, ionisation, and fragmentation logic
- Mass Spectrometry for Proteomics (PMC) — instrument types, tandem MS methods, and PTM analysis strategies
- An Introduction to MS-Based Proteomics, Journal of Proteome Research — accessible overview of sample prep, data acquisition, and analysis
- The Hunt Lab Guide to De Novo Peptide Sequence Analysis — b/y ion series, MS/MS interpretation, and free fragment calculator tools
- A Beginner’s Guide to MS-Based Proteomics, Portland Press — ESI mechanics, Orbitrap vs TOF, and sample preparation principles
- Peptide purity standards: HPLC, mass spec, and what >98% really means — HPLC area percent limitations and why MS is an orthogonal identity check
- How to Verify Peptide Identity: Mass Spectrometry for Beginners — charge state calculation, common mass shifts, and COA field requirements
- Mass spectrometry for peptides: verifying identity and molecular weight — MALDI-TOF vs ESI comparison, tolerance standards, and LC-MS for purity
Peptilab pages referenced for Canadian procurement and COA context: research peptides Canada, batch QC documentation, peptide sequence verification.